IDMP: The Latest in Identification of Medicinal Products
While some deadlines for economic operator registration have already passed, the rollout of the device registration module will necessitate a proactive data management strategy.
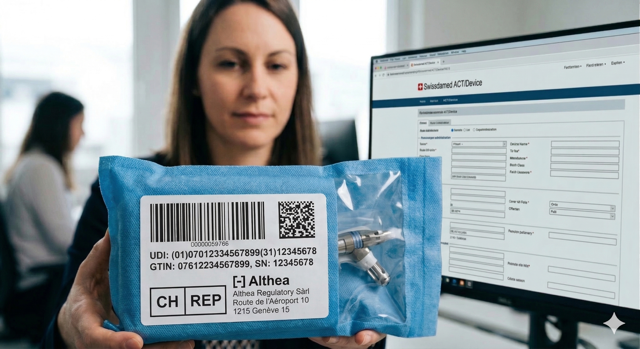
The Device Registration (UDI) Module became available for voluntary registration in August 2025. This allows companies to begin uploading data for medical devices, IVDs, and systems/procedure packs. While currently optional, July 1, 2026, marks the start of the mandatory registration phase. From this date, any new device placed on the Swiss market must be registered before its launch. For products already on the market, a transitional period exists until December 31, 2026, by which point all devices must be fully registered in Swissdamed.
Companies should not view Swissdamed as a mere clerical task; it is a legal prerequisite for market access. Ensuring that your data is accurate and synchronized with your technical documentation is essential to avoid delays in product placement or potential enforcement actions from Swissmedic.
CH-REP
Central to this new Swiss framework is the role of the Swiss Authorised Representative (CH-REP). For any manufacturer based outside of Switzerland, the appointment of a CH-REP is not an optional convenience but a mandatory legal anchor. The CH-REP acts as the formal liaison between the manufacturer and the Swiss market surveillance authorities. Their responsibilities go beyond simply holding a certificate; they are legally liable for the device’s compliance. This includes verifying that the Declaration of Conformity and technical documentation have been drawn up, and ensuring that the manufacturer has fulfilled their registration obligations.
From a practical standpoint, the CH-REP must be integrated into the manufacturer’s quality management system. They must have permanent access to the technical documentation and be prepared to respond to inquiries from Swissmedic within short timeframes. For many companies, the transition periods for appointing a CH-REP have already lapsed for higher-risk classes, but for certain In Vitro Diagnostics (IVDs) and legacy devices, the window for compliance is narrowing. It is advisable to review current mandates to ensure the CH-REP is not just a name on a label, but a functional partner capable of managing the regulatory burden.
PRRC
Closely linked to the CH-REP is the requirement for a Person Responsible for Regulatory Compliance (PRRC). Under the Swiss Medical Devices Ordinance (MedDO) and the Ordinance on In Vitro Diagnostic Medical Devices (IvDO), both the manufacturer and the CH-REP must have a PRRC at their disposal. The PRRC’s mandate is clear: they must ensure that the conformity of the devices is appropriately checked, that the technical documentation and the EU declaration of conformity are kept up to date, and that post-market surveillance obligations are met.
The challenge for many organizations lies in the qualifications and the availability of the PRRC. This individual must possess the requisite professional knowledge, usually evidenced by a degree in a relevant scientific or legal discipline and several years of professional experience in regulatory affairs or quality management related to medical devices. In the Swiss context, the PRRC of the CH-REP ensures that the representative can fulfill their oversight duties effectively. This dual-layer of accountability—at the manufacturing level and the representative level—is designed to close gaps in market safety.
For companies operating in Switzerland, the immediate priority should be a comprehensive “gap analysis” of their Swiss-specific documentation. It is vital to ensure that labeling, Instructions for Use (IFU), and symbols correctly identify the CH-REP. Furthermore, manufacturers should begin preparing their data for Swissdamed registration now, rather than waiting for the platform’s final deadlines.
The Swiss market remains a lucrative and stable environment for medical innovation, but the cost of entry is now defined by administrative diligence. Compliance is not a static achievement but a continuous process of alignment with Swissmedic’s evolving digital and legal requirements. By securing a robust CH-REP partnership and clarifying the roles of the PRRC, manufacturers can mitigate the risks of market exclusion and focus on the primary goal: providing safe and effective technology to the Swiss market. More news here : https://www.swissmedic.ch/swissmedic/en/home/medical-devices/medizinprodukte-datenbank.html
As the pharmaceutical landscape continues to evolve, the need for standardized identification and management of medicinal products has never been more critical. The Identification of Medicinal Products (IDMP) standards, established by the International Organization for Standardization (ISO) and supported by regulatory bodies like the European Medicines Agency (EMA), are crucial for ensuring global consistency, safety, and efficiency in drug development and regulation. As of 2024, these standards are more relevant than ever for life sciences professionals.
What is IDMP?
IDMP encompasses a suite of five ISO standards (ISO 11615, ISO 11616, ISO 11238, ISO 11239, and ISO 11240) aimed at harmonizing the identification and exchange of medicinal product information. These standards are designed to facilitate the clear and consistent communication of product information across regulatory agencies and healthcare systems worldwide.
Key IDMP Standards
- ISO 11615: Defines the identification of medicinal products, including both authorized and investigational drugs.
- ISO 11616: Pertains to the identification of pharmaceutical products, specifying dose form and administration details.
- ISO 11238: Relates to the identification of substances, including active ingredients and excipients.
- ISO 11239: Addresses pharmaceutical dose forms, units of presentation, routes of administration, and packaging.
- ISO 11240: Standardizes units of measurement used in medicinal product descriptions.
Benefits of IDMP Implementation
Regulatory Compliance and Global Harmonization
The EMA and other regulatory bodies have mandated the adoption of IDMP standards, making compliance essential for market access. These standards ensure that medicinal product data is consistent across different jurisdictions, streamlining regulatory submissions and approvals. For example, the EMA’s implementation of the SPOR (Substances, Products, Organizations, and Referentials) data management services is directly aligned with IDMP requirements.
Enhanced Pharmacovigilance and Patient Safety
IDMP standards improve pharmacovigilance by enabling more accurate tracking and reporting of adverse drug reactions. Standardized product information facilitates quicker identification of safety issues, supporting better risk management and enhancing patient safety.
Operational Efficiency and Data Integrity
Implementing IDMP can significantly improve operational efficiency within pharmaceutical companies. Standardized data formats reduce the risk of errors and discrepancies, leading to more reliable data management and interoperability between different systems and stakeholders.
Recent Developments and Challenges
EMA’s SPOR Initiative
The EMA’s SPOR initiative, which aligns with IDMP, has seen significant progress. The SPOR platform enables the efficient management of substance, product, organization, and referential data, providing a robust foundation for IDMP compliance. The ongoing updates to SPOR services, including enhanced data validation and integration capabilities, are crucial for maintaining compliance and operational efficiency in 2024.
Implementation Challenges
Despite the clear benefits, implementing IDMP is not without challenges. Data harmonization across legacy systems, integrating new IT solutions, and ensuring staff are trained on new processes require significant resources and strategic planning. Companies must develop comprehensive implementation plans, often requiring expert guidance to navigate these complexities effectively.
How Helix Life Sciences Can Support Your IDMP Journey
At Helix Life Sciences, we understand the intricacies of IDMP implementation and the challenges it presents. Our team of experts offers tailored consulting services to help pharmaceutical companies achieve compliance with the latest IDMP standards. We provide strategic support, from data harmonization and system integration to staff training and regulatory submission preparation.
For more information on how we can assist with your IDMP needs, visit our website or contact us directly. Helix Life Sciences is committed to helping you navigate the complexities of IDMP, ensuring your products meet the highest standards of regulatory compliance and patient safety.
Related Posts